Living With Pompe Disease: Facts, Treatment, and Support
If you or someone you love has been told they might have Pompe disease, you probably have a lot of questions. It’s a rare condition, and many people have never heard of it until it touches their own family. Accessia Health is here to provide resources and financial support for those with Pompe disease through our patient program.
What Is Pompe Disease?
Pompe disease, also called glycogen storage disease type II or acid maltase deficiency, is an inherited metabolic disorder. Harmful amounts of glycogen, a stored form of sugar, build up inside the cells’ recycling compartments, called lysosomes. The cause is a shortage of an enzyme called GAA. Mutations in the GAA gene reduce or completely eliminate it, and researchers have identified up to 300 different mutations that can cause the disease. Without enough GAA, glycogen accumulates and gradually damages muscles, including the heart and the muscles used for breathing.
How Common Is Pompe Disease?
Pompe disease is rare. It is estimated to affect about 1 in 40,000 people in the U.S. Rates vary among ethnic groups.
Newborn screening is changing the picture. Screening data from Japan from 2013 to 2020 showed an incidence of about 1 in 37,000. Researchers expect screening to give us more accurate numbers over time.
The Passing Down of Pompe Disease
Pompe disease is inherited in an autosomal recessive pattern. A child must receive one changed copy of the gene from each parent. Parents who each carry one copy typically show no signs of the disease themselves. That’s why Pompe can seem to appear “out of nowhere” in a family.
Doctors usually describe two main forms:
- Infantile-onset Pompe disease starts in the first months of life. It is the result of a severe GAA deficiency. Without treatment, the hearts of affected babies progressively thicken and enlarge.
- Late-onset Pompe disease is caused by a less severe GAA deficiency. Symptoms can appear as early as the first year of life or as late as the sixth decade. The heart is rarely involved to the extent seen in infants.
Signs and Symptoms of Pompe disease
Symptoms depend on the age of onset and how much working enzyme the body has. In general, the disease is progressive, meaning symptoms worsen over time.
In infantile-onset Pompe, signs can include a very weak or “floppy” feel, trouble feeding, breathing difficulties, and heart problems. Many infants also have an enlarged tongue.
In older children and adults with late-onset Pompe, the main feature is muscle weakness that can progress to weakness in the muscles used for breathing. The condition resembles limb-girdle muscular dystrophy. In everyday life, this may look like:
- Trouble climbing stairs or getting up from a chair
- Frequent tiredness or difficulty keeping up with others
- Shortness of breath, especially when lying down
- Frequent stumbling or falls
These everyday examples are general descriptions, not drawn from a specific source. A doctor can tell you whether they point to Pompe disease or something else.
Why Answers Can Take Time
Pompe disease is hard to identify because many of its symptoms overlap with other conditions, such as multiple sclerosis and muscular dystrophy. Many people see several doctors before getting answers.
Diagnosis can be confirmed by testing for common genetic mutations or by measuring GAA enzyme activity in a blood sample. The diagnosis is ultimately confirmed with enzyme assays and genetic testing. Once a diagnosis is made, testing of family members and a visit with a genetic specialist are recommended.
Newborn screening has made a real difference. Pompe disease was added to the national Recommended Uniform Screening Panel in 2015, and as of a 2023 report, 38 states were screening newborns for it. A positive screen doesn’t always mean a baby has the disease. Not all infants with low GAA activity on screening will have Pompe disease, so follow-up testing matters.
Pompe Disease Treatment & Daily Management
There is currently no cure for Pompe disease. Treatment centers on enzyme replacement and supportive care.
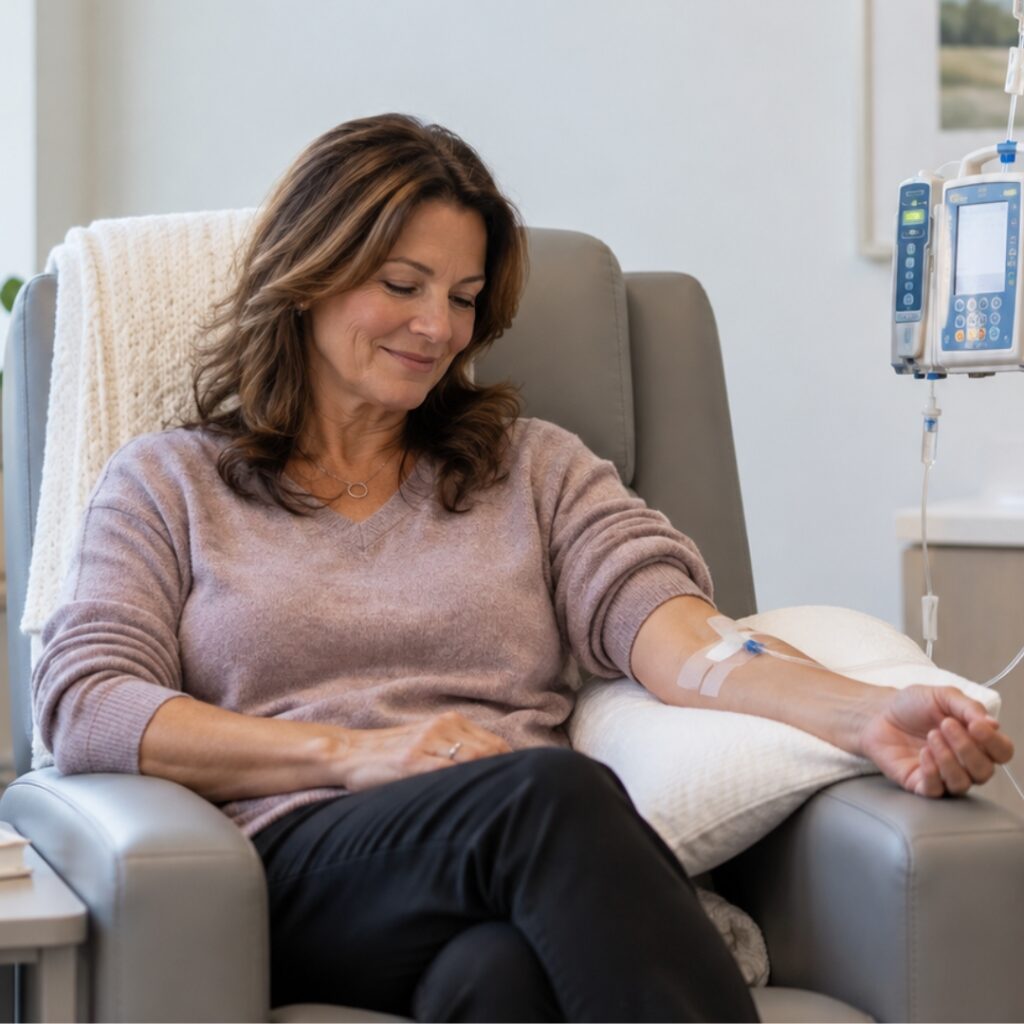
- Enzyme replacement therapy (ERT) is the main treatment. It involves regular intravenous infusions of a lab-made version of the missing GAA enzyme to help clear glycogen buildup in cells. In clinical trials with infantile-onset patients, ERT decreased heart size, maintained normal heart function, improved muscle function and strength, and reduced glycogen accumulation. If infantile Pompe disease is diagnosed, ERT should be started as soon as possible.
- Supportive care matters too. It can include respiratory support, mobility support, physical therapy, and dietary care. Physical therapy has an important role alongside ERT.
- Research continues. Several companies are pursuing gene therapies for Pompe. Glycogen reduction approaches are also in development.
Families should know that treatment is not curative, and problems can’t always be prevented. Even so, early diagnosis and intervention are widely regarded as key to better outcomes. Living with Pompe is easier with a knowledgeable team and a strong support system.
Accessia Health’s Pompe Patient Program
Rare disease is hard in part because it can feel isolating. Appointments, infusions, insurance questions, and day-to-day changes in strength and energy add up, for patients and caregivers alike.
That’s why Accessia Health has created a patient program for people living with Pompe disease. If you have Pompe, or you care for someone who does, we encourage you to apply.
Who should apply?
- People diagnosed with Pompe disease, whether infantile-onset or late-onset
- Parents and caregivers of a child with Pompe
- Anyone with a family member who has Pompe disease
[Learn More About Our Pompe Patient Program]
This post is for general education and is not medical advice. Please talk with a qualified healthcare provider about any health concerns.